From gene-targeted approaches in Dravet syndrome to positive phase 3 data in focal and generalized seizures, selected presentations at the 16th European Epilepsy Congress highlighted a rapidly evolving epilepsy treatment landscape. Beyond reducing seizure frequency, several studies also explored whether emerging therapies can improve development, behavior, quality of life, and meaningful periods of seizure freedom.
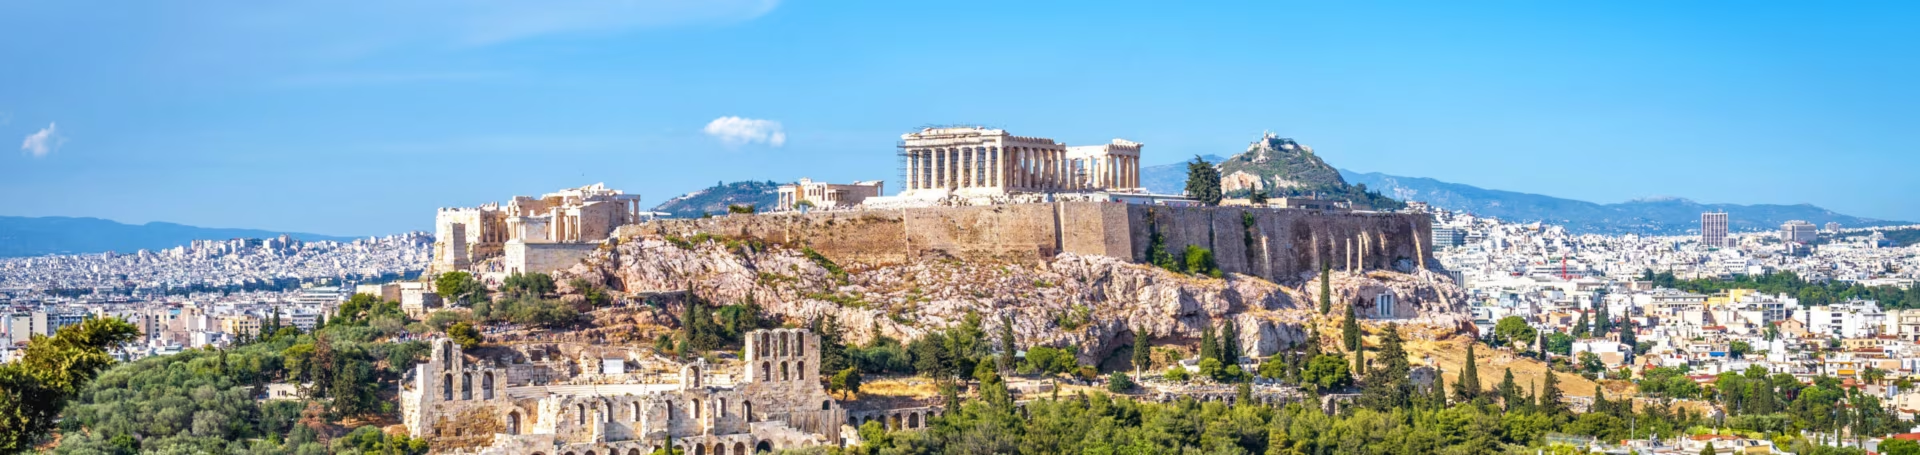
The 16th European Epilepsy Congress, organized by the International League Against Epilepsy, took place in Athens, Greece, from September 5–9, 2026. Across the congress, an emerging theme was the move toward treatments designed around the underlying biology of epilepsy, particularly for developmental and epileptic encephalopathies, alongside new antiseizure medications for more common drug-resistant epilepsies.
Disease-modifying approaches show promise in Dravet syndrome
Zorevunersen shows potential beyond seizure control
Prof. J. Helen Cross (UCL NIHR BRC Great Ormond Street Institute of Child Health, London, UK)Abstract 1101 discussed long-term findings for zorevunersen, an investigational antisense oligonucleotide designed to increase NaV1.1 protein expression from the unaffected copy of the SCN1A gene.1 Among 81 patients who received zorevunersen in phase 1/2a studies, 75 entered the open-label extension studies. Durable reductions in seizure frequency were accompanied by improvements in quality of life and overall clinical status.
Among seven patients who had received two or three 70 mg loading doses, mean quality-of-life scores improved by 32.1% at Month 20, while 94.7% of evaluable patients showed improvement in overall clinical status at Month 36.1
Developmental outcomes add to the zorevunersen findings
Prof. Andreas Brunklaus Abstract 1131 (University of Glasgow, Glasgow, UK) presented a matching-adjusted indirect comparison between patients receiving phase 3-like zorevunersen dosing and patients from the BUTTERFLY natural history study.2 At Week 100/104, estimated mean improvements across five Vineland Adaptive Behavior Scales, Third Edition, subdomains ranged from 9.8 to 14.4 points with zorevunersen, compared with changes ranging from −2.6 to 2.8 points in the natural history group.
Updated longer-term findings presented around the congress also suggested that improvements in expressive and receptive communication, interpersonal relationships, personal skills, and coping skills were sustained through 4 years of treatment, whereas patients in the natural history cohort showed relatively little change over a comparable period.3
These indirect comparisons cannot establish disease modification on their own, but the divergence in developmental trajectories provides an important rationale for the ongoing phase 3 EMPEROR study (NCT06872125).
Gene regulation therapy offers another approach to Dravet syndrome
Prof. Ingrid E. Scheffer Abstract 1378 (Epilepsy Research Centre, The University of Melbourne and Austin Health, Melbourne, Australia) discussed updated findings from the POLARIS program evaluating ETX101.4 ETX101 is an investigational adeno-associated virus serotype 9-based gene regulation therapy designed to increase endogenous SCN1A expression selectively in inhibitory interneurons following a single intracerebroventricular administration.
Updated data presented at the congress extended beyond those contained in the original abstract. Across Week 5 to Week 52, or the latest available assessment, median monthly countable seizure frequency fell by approximately 76% among patients receiving dose level 3 (n=5) and 60% among those receiving dose level 4 (n=9). Among patients with 52 weeks of follow-up, reductions at Month 12 reached approximately 79% and 89%, respectively.5
Neurodevelopmental findings were also notable. Children treated before 2 years of age demonstrated continued cognitive gains, with trajectories increasingly diverging from developmental stagnation seen in the ENVISION natural history study over follow-up of up to 76 weeks. Improvements were also reported across communication, motor skills, socialization, and daily living skills.5
With up to 117 weeks of safety follow-up, no treatment- or procedure-related serious adverse events had been reported. However, these findings are derived from small, open-label cohorts, and confirmation in the ongoing pivotal ENDEAVOR program will be important.
Together, the zorevunersen and ETX101 programs demonstrate how Dravet syndrome is becoming an important test case for disease-modifying epilepsy treatment: one approach modulates RNA processing to increase functional protein expression, while the other uses one-time gene regulation to address the same underlying SCN1A deficit.
Phase 3 fenfluramine data in CDKL5 deficiency disorder
Fenfluramine reduces motor seizure frequency
Prof. Renzo Guerrini Abstract 803 (Meyer Children’s Hospital IRCCS and University of Florence, Florence, Italy) presented results from a randomized, double-blind, placebo-controlled phase 3 study (NCT05064878) of fenfluramine in children and adults aged 1–35 years with CDKL5 deficiency disorder.6
In the modified intention-to-treat population of 86 patients, median countable motor seizure frequency decreased by 47.6% with fenfluramine compared with 2.8% with placebo (p<0.001). A ≥50% reduction was achieved by 45.2% and 4.5% of patients, respectively.6
Clinically meaningful improvement on the investigator-assessed Clinical Global Impression-Improvement scale was also more common with fenfluramine than placebo, at 38.1% versus 6.8%. No new safety signals, valvular heart disease, pulmonary arterial hypertension, or deaths were reported during the randomized period.
Seizure-free days provide a different measure of treatment benefit
Dr Ángel Aledo-Serrano Abstract 606 (Blua Sanitas Valdebebas Hospital, Madrid, Spain) explored the same trial through a different endpoint: seizure-free days.7 Patients receiving fenfluramine experienced a median increase of 6.4 countable motor seizure-free days per month from Baseline, compared with 0.1 days with placebo. The median longest period without countable motor seizures was 8.5 days with fenfluramine compared with 3.0 days with placebo.
During treatment, 71.4% of patients receiving fenfluramine experienced more than seven countable motor seizure-free days per month, compared with 31.8% receiving placebo. More than 14 seizure-free days per month were achieved by 59.5% and 6.8% of patients, respectively.7
Although post hoc, these findings provide a more tangible perspective on treatment benefit than percentage seizure reduction alone.
Fenfluramine extends the time between seizure events
Dr Antonio Gil-Nagel Abstract 698 (Hospital Ruber Internacional de Madrid, Madrid, Spain) presented a complementary time-to-event analysis.8 The median time taken for patients to accumulate the same number of countable motor seizures experienced during Baseline was 55 days with fenfluramine compared with 29 days with placebo. For all seizures, the corresponding times were 51 and 29 days, respectively.
Collectively, these analyses strengthen the efficacy signal from the primary phase 3 study while also illustrating a broader change in how outcomes in severe developmental epilepsies are being considered. Longer intervals between seizures and additional seizure-free days may provide benefits to patients and families that are not fully captured by conventional responder thresholds.
Looking beyond seizures in Lennox-Gastaut syndrome
Fenfluramine may influence everyday executive function
Patrick Healy Abstract 1941 (UCB, Morrisville, NC, USA) discussed a post hoc analysis evaluating everyday executive function in pediatric patients aged 6–18 years who participated in the phase 3 fenfluramine study (NCT03355209).9
Clinically meaningful improvement in the Global Executive Composite score was observed in 27.3% of patients receiving fenfluramine 0.7 mg/kg/day, compared with 16.7% receiving 0.2 mg/kg/day and 14.3% receiving placebo.
The findings should be considered exploratory, particularly as the analysis was post hoc and involved a subset of the original trial population. Nevertheless, assessing executive function alongside seizures is relevant in Lennox-Gastaut syndrome, where cognitive and behavioral difficulties contribute substantially to overall disease burden.
Cannabidiol studies explore outcomes beyond seizure frequency
Neuropsychiatric outcomes in tuberous sclerosis complex
Joanne Stevens Abstract 394 (Jazz Pharmaceuticals, Inc., Philadelphia, PA, USA) presented a 6-month intermediate analysis from the ongoing EpiCom study (NCT05864846), which is examining tuberous sclerosis complex-associated neuropsychiatric disorders following initiation of adjunctive cannabidiol.10
Among 62 participants with at least one post-Baseline assessment, the median severity score for the most problematic behavior was 9.0 out of 10 at Baseline. At Week 26, the median change was −2.0 points.
Improvements were also observed across measures of eating and sleep, overactivity and impulsivity, irritability, and hyperactive noncompliance. As EpiCom is open label, these changes cannot be attributed definitively to cannabidiol; however, the study highlights the importance of considering tuberous sclerosis complex-associated neuropsychiatric disorders alongside seizure control.
Long-term cannabidiol data in adults with Lennox-Gastaut syndrome
Kishan Vyas Abstract 415 (Jazz Pharmaceuticals UK Ltd., London, UK) focused on a different evidence gap: cannabidiol treatment in adults with Lennox-Gastaut syndrome.11 In pooled phase 3 data, 74 adults receiving cannabidiol experienced median reductions of 33.9% in drop seizures and 31.6% in total seizures by the end of the randomized studies.
Among patients continuing into the open-label extension, reductions at the end of available follow-up were 68.8% and 64.6%, respectively. One- and 3-year retention rates in the extension study were 80% and 70%, respectively.11
As this was a descriptive post hoc analysis and was not designed to provide a new statistical comparison with placebo, the results should be interpreted accordingly, but they provide useful longer-term evidence in an adult population that is often underrepresented in Lennox-Gastaut syndrome studies.
New treatment options for focal and generalized seizures
Azetukalner meets phase 3 endpoints in focal onset seizures
Prof. Jacqueline French Abstract 726 (NYU Grossman School of Medicine and NYU Langone Health, New York, NY, USA) presented results from the phase 3 X-TOLE2 study (NCT05614063) of azetukalner, a novel KV7 potassium channel opener, in adults with focal onset seizures.12 The study randomized 380 adults receiving one to three background antiseizure medications, with participants having previously tried and discontinued a median of five treatments.
Final efficacy data showed a 53.2% median reduction from Baseline in monthly focal onset seizure frequency with azetukalner 25 mg, compared with 34.5% with 15 mg and 10.4% with placebo. Both doses met the primary endpoint, with the 25 mg dose producing a placebo-adjusted reduction of 42.7%.12,13
Azetukalner was administered once daily without titration and was generally well tolerated, with no new safety signals identified.
Longer-term X-TOLE extension data have also suggested sustained benefit. Among patients remaining in the study at 48 months, almost 40% had experienced at least one continuous 12-month period of seizure freedom and approximately one-quarter had achieved at least 24 months.13 These open-label extension findings are subject to survivor and selection effects but provide encouraging evidence of durability.
Cenobamate shows efficacy in primary generalized tonic-clonic seizures
Dr David G. VosslerAbstract 836 (University of Washington School of Medicine, Seattle, WA, USA) discussed a phase 3 trial (NCT03678753) extending the evidence for cenobamate into primary generalized tonic-clonic seizures.14
Among 168 patients included in the analysis, median 28-day seizure frequency decreased by 71.9% with cenobamate compared with 39.6% with placebo (p=0.003). During maintenance treatment, 71.1% of patients receiving cenobamate achieved a ≥50% response compared with 51.3% receiving placebo.
Notably, 43.4% of patients receiving cenobamate were seizure free during the maintenance phase, compared with 20.5% receiving placebo (p=0.002). Treatment-emergent adverse events occurred in 60.0% and 53.0% of patients, respectively, with somnolence and fatigue the most common events with cenobamate. No treatment-related serious adverse events, deaths, or cases of drug reaction with eosinophilia and systemic symptoms were reported.14
Vormatrigine produces rapid seizure reductions in an open-label study
Dr Ángel Aledo-SerranoAbstract 913 (Blua Sanitas Valdebebas Hospital, Madrid, Spain) presented findings from RADIANT (NCT06908356), an open-label phase 2 study of vormatrigine in adults with treatment-resistant epilepsy.15
Among 62 patients with focal onset seizures, median seizure frequency decreased by 54% overall. A >50% reduction was observed in 58% of patients by Week 1 and 61% by Week 8, while more than 11% achieved seizure freedom and approximately one-third experienced a consecutive 28-day seizure-free period.
The uncontrolled design of RADIANT makes these findings preliminary, and subsequent data provide important context. In June 2026, the randomized phase 2/3 POWER1 study of vormatrigine in highly treatment-resistant focal onset seizures did not meet its primary success measure, although the ≥50% responder secondary measure was met and seizure reduction appeared greater during the later 30 mg dosing period.16
The contrast between the encouraging RADIANT signal and the POWER1 primary endpoint miss illustrates the importance of controlled studies when assessing promising early-stage epilepsy treatments.
Precision therapies target genetically defined epilepsies
Relutrigine targets sodium channel-related developmental epilepsies
Dr Antonio Gil-Nagel Abstract 907 (Hospital Ruber Internacional de Madrid, Madrid, Spain) discussed the EMBOLD study (NCT05818553), which evaluated relutrigine in pediatric patients with SCN2A– and SCN8A-related developmental and epileptic encephalopathies.17 Relutrigine is designed to preferentially modulate sodium channels in hyperexcitable neuronal states.
Across the randomized study, relutrigine was associated with a 53% placebo-adjusted reduction in motor seizure frequency over 16 weeks (n=51; p<0.0002), alongside a 66% increase in seizure-free days from Baseline (p=0.034).
Statistically significant improvements were also reported in clinician- and caregiver-assessed measures of alertness, communication, behavior, and seizure severity. The data monitoring committee recommended stopping EMBOLD early for efficacy.17
Although the findings suggest benefits beyond seizure reduction, longer-term evidence will be important in establishing whether these translate into modification of the developmental trajectory associated with SCN2A– and SCN8A-related disease.
Novel approaches to treatment-resistant epilepsy
Interneuron cell therapy aims to restore inhibitory signaling
Dr Eduardo DunayevichAbstract 826 (Neurona Therapeutics, South San Francisco, CA, USA) presented an update on NRTX-1001, an investigational human gamma-aminobutyric acid-producing interneuron cell therapy being evaluated in adults with drug-resistant mesial temporal lobe epilepsy.18
The treatment involves a one-time transplantation of human pluripotent stem cell-derived interneurons into the hippocampus, with the aim of restoring inhibitory signaling within seizure-generating circuitry.
Among nine patients with mesial temporal sclerosis treated at the lower dose, median disabling seizure frequency decreased by 89% during Months 7–12, with 78% achieving a >50% reduction. In nine patients receiving the higher dose, the median reduction was 78% during the most recently completed 6-month assessment period, with 89% achieving a >50% response.18
Importantly, seizure control persisted after completion of immunosuppression, and no overall trend toward cognitive decline was observed. At 18 months, six of eight evaluable patients reported a clinically meaningful improvement in quality of life.
These remain early, open-label data, but a cell-based strategy designed to restore inhibitory circuitry represents a substantially different approach from chronic pharmacological seizure suppression.
Treating an ultra-rare metabolic epilepsy
The MEND-PNPO study evaluates pharmaceutical-grade pyridoxal 5′-phosphate
Dr Phillip L. Pearl Abstract 566 (Harvard Medical School, Boston, MA, USA) presented the design and rationale of the phase 3 MEND-PNPO study (NCT04706013), which is evaluating MC-1, a pharmaceutical-grade formulation of pyridoxal 5′-phosphate, in pyridox(am)ine 5′-phosphate oxidase deficiency.19
Pyridox(am)ine 5′-phosphate oxidase deficiency is an ultra-rare neurometabolic condition in which impaired vitamin B6 metabolism can cause severe neonatal seizures. Although pyridoxal 5′-phosphate can be effective, currently available nonpharmaceutical preparations may vary in dose and purity.
MEND-PNPO is a 52-week, open-label, externally controlled study designed to enroll approximately 10 patients. Updated congress reporting indicated that nine patients had been enrolled and three had completed 1 year of treatment.20
The presentation primarily concerned study design, pharmacokinetics, and Baseline characteristics rather than definitive phase 3 efficacy results. The safety and effectiveness of MC-1 therefore remain to be established.
Could epilepsy eventually be prevented?
Janus kinase pathway inhibition offers an intriguing signal
Dr Katharina Schuler Abstract 1301 (University Hospital Zurich, Department of Neurology, Zurich, Switzerland) investigated whether inhibition of the Janus kinase/signal transducer and activator of transcription pathway could influence the development of epilepsy.21
Using international real-world data, investigators propensity matched 5,027 patients with rheumatoid arthritis initiating tofacitinib with 5,027 patients initiating tumor necrosis factor-α inhibitors; none had a history of seizures or epilepsy.
Treatment with tofacitinib was associated with a 58% lower risk of subsequently developing seizures or epilepsy (hazard ratio 0.42; 95% confidence interval 0.22–0.84). A sensitivity analysis incorporating other approved Janus kinase/signal transducer and activator of transcription inhibitors also found a reduced risk.21
These observational data cannot demonstrate that pathway inhibition prevents epilepsy, and residual confounding remains possible. Nevertheless, the findings provide an interesting human signal for a pathway previously implicated in epileptogenesis and could support prospective investigation of seizure prevention in appropriately selected high-risk populations.
Moving beyond seizure frequency
Taken together, selected findings from the European Epilepsy Congress 2026 highlighted two parallel shifts in epilepsy research.
For focal and generalized epilepsies, treatments such as azetukalner and cenobamate are seeking to improve seizure control in populations that continue to experience seizures despite existing therapies. At the same time, treatments for genetically defined developmental epilepsies are becoming increasingly precise, with antisense oligonucleotides, gene regulation, and selective sodium channel modulators designed around the molecular mechanisms responsible for disease.
The definition of treatment success may also be changing. Across the zorevunersen, ETX101, relutrigine, fenfluramine, and cannabidiol programs, investigators evaluated outcomes including adaptive development, executive function, behavior, quality of life, seizure-free days, and the duration of seizure-free periods.
Many of these findings remain exploratory, post hoc, open label, or based on relatively small populations, and robust controlled data are still required. However, the congress provided a clear indication of the direction of the field: toward therapies that are increasingly tailored to disease biology and judged not only by how much they reduce seizures, but also by whether they meaningfully alter patients’ everyday lives.
References
- Knupp K, Sullivan J, Perry MS, et al. Zorevunersen demonstrates disease-modifying potential in patients with Dravet syndrome with improvements in seizure burden, quality of life, and overall clinical status. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 1101.
- Brunklaus A, Cross JH, Knupp K, et al. Zorevunersen demonstrates potential as a disease-modifying therapy in patients with Dravet syndrome: A matching-adjusted indirect comparison between natural history and patients receiving zorevunersen. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 1131.
- Meglio M. Investigational zorevunersen linked to adaptive gains in Dravet syndrome. NeurologyLive. 2026. Available at: www.neurologylive.com/view/investigational-zorevunersen-linked-adaptive-gains-dravet-syndrome (accessed September 10, 2026).
- Scheffer IE, Howell KB, Sullivan J, et al. Interim safety and preliminary efficacy of ETX101, a gene regulation therapy for SCN1A+ Dravet syndrome: Results from the POLARIS phase 1/2 program. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 1378.
- Encoded Therapeutics. Encoded Therapeutics presents updated data from POLARIS phase 1/2 trials of ETX101 gene therapy in Dravet syndrome at the 16th European Epilepsy Congress [Press release]. September 8, 2026. Available at: www.businesswire.com/news/home/20260908732226/en/Encoded-Therapeutics-Presents-Updated-Data-from-POLARIS-Phase-12-Trials-of-ETX101-Gene-Therapy-in-Dravet-Syndrome-at-the-16th-European-Epilepsy-Congress (accessed September 10, 2026).
- Specchio N, Marsh E, Devinsky O, et al. Fenfluramine in CDKL5 deficiency disorder: Primary efficacy and safety results from a phase 3, randomized, double-blind, placebo-controlled study. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 803.
- Aledo-Serrano Á, Demarest S, Olson H, et al. Examining seizure-free days with fenfluramine in children and adults with CDKL5 deficiency disorder: A post hoc analysis of a phase 3, randomized, double-blind, placebo-controlled trial. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 606.
- Marsh E, Gil-Nagel A, Aledo-Serrano Á, et al. Seizure control with fenfluramine versus placebo in children and adults with CDKL5 deficiency disorder: A post hoc time-to-event analysis of data from a phase 3, randomized, placebo-controlled trial. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 698.
- Breuillard D, Knupp KG, Healy P, et al. Fenfluramine-associated changes in everyday executive function in pediatric patients with Lennox-Gastaut syndrome: Post hoc analysis of a phase 3 trial. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 1941.
- van Eeghen A, Wilson SM, Roszkowski S, et al. Tuberous sclerosis complex-associated neuropsychiatric disorders outcomes following adjunctive cannabidiol treatment: 6-month intermediate analysis of the EpiCom trial. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 394.
- Vyas K, Greco T, Harrison M, Lattanzi S. Efficacy and safety of cannabidiol in adult patients with Lennox-Gastaut syndrome: A post hoc analysis of phase 3 and open-label extension data. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 415.
- French J, LaFrance WC Jr, Ryvlin P, et al. Results from the phase 3 X-TOLE2 study evaluating azetukalner, a novel, potent KV7 channel opener, in adults with focal onset seizures. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 726.
- Xenon Pharmaceuticals. Xenon announces positive topline data from phase 3 X-TOLE2 study of azetukalner in focal onset seizures [Press release]. March 9, 2026. Available at: investor.xenon-pharma.com/news-releases/news-release-details/xenon-announces-positive-topline-data-phase-3-x-tole2-study (accessed September 10, 2026).
- Vossler D, Shih E, Bar M, et al. Efficacy and safety of adjunctive cenobamate for the treatment of primary generalized tonic-clonic seizures in adults and adolescents. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 836.
- Hansen K, Mohsen B, Epstein N, et al. Vormatrigine rapidly reduces seizures in adults with treatment-resistant epilepsy: Results from the RADIANT study. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 913.
- Praxis Precision Medicines. Praxis Precision Medicines provides vormatrigine program update [Press release]. June 1, 2026. Available at: praxismedicines.gcs-web.com/news-releases/news-release-details/praxis-precision-medicines-provides-vormatrigine-program-update (accessed September 10, 2026).
- Kamireddy S, Frizzo S, Spar B, et al. Efficacy and safety of relutrigine in pediatric participants with SCN2A- and SCN8A-related developmental and epileptic encephalopathies: Results from the EMBOLD study. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 907.
- Dunayevich E, Bershteyn M, Bulfone A, et al. Clinical update: NRTX-1001 GABAergic interneuron cell therapy for drug-resistant focal epilepsy. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 826.
- Pearl P, Cole L, Stewart N, et al. Pharmaceutical grade oral pyridoxal-5-phosphate for the treatment of patients with PNPO deficiency: MEND-PNPO study design and rationale. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 566.
- ScanX. Medicure presents MEND-PNPO phase 3 study data at European Epilepsy Congress. 2026. Available at: scanx.trade/stock-market-news/companies/medicure-presents-mend-pnpo-phase-3-study-data-european-epilepsy/50452892 (accessed September 10, 2026).
- Schuler K, Ferreira-Atuesta C, Schubert KM, et al. Antiepileptogenic effects of JAK/STAT pathway inhibition in humans: A large-scale matched cohort study. Presented at: 16th European Epilepsy Congress, Athens, Greece, September 5–9, 2026. Abstr 1301.
More content in epilepsy
Cite: European Epilepsy Congress 2026: Disease-modifying and precision therapies reshape the treatment landscape. touchNEUROLOGY. 18 September 2026.
Editor: Katey Gabrysch, Editorial Director.
Disclosures: The content was developed and edited by human editors. No fees or funding were associated with its publication. touchNEUROLOGY utilize AI as an editorial tool (ChatGPT (GPT-4o) [Large language model]. https://chat.openai.com/chat).
This content has been developed independently by Touch Medical Media for touchNEUROLOGY. It is not affiliated with European Epilepsy Congress 2026. Views expressed are the speaker’s own and do not necessarily reflect the views of Touch Medical Media.
SIGN UP to touchNEUROLOGY!
Join our global community today for access to thousands of peer-reviewed articles, expert insights, and learn-on-the-go education across 150+ specialties, plus concise email updates and newsletters so you never miss out.